
European Union Member States are continuing work on Friday, 11 September 2026, on changes to the rules for medical devices and in vitro diagnostic tests. The file is being examined by the Council of the EU’s Working Party on Pharmaceuticals and Medical Devices. This is a technical stage of the legislative procedure, not the adoption of a new regulation and not the entry into force of a new system.
On 16 December 2025 the European Commission presented proposal COM(2025) 1023, which would amend Regulation (EU) 2017/745 on medical devices (the MDR) and Regulation (EU) 2017/746 on in vitro diagnostic medical devices (the IVDR). The MDR became applicable, in the main, in 2021 and the IVDR in 2022. The Commission’s stated aim is to simplify certain procedures and reduce administrative burdens while maintaining a high level of patient protection. The proposal does not yet replace the legal framework now in force.
The two regulations cover a very wide range of products used in hospitals, laboratories and clinics: from implants and pacemakers to imaging equipment, surgical instruments and laboratory tests on biological samples. The Union introduced them to strengthen safety, oversight and traceability. Implementation has, however, also exposed heavy procedures for certification, documentation and surveillance, including for small and medium-sized firms, which make up a large part of Europe’s medical-technology industry.
Conformity assessment is a central issue in the talks. For higher-risk devices, manufacturers generally need a notified body before a product can be placed on the European market. The capacity of those bodies and the time taken by procedures have become a point of tension: a system that is too slow or unpredictable can delay new devices or contribute to products being withdrawn, especially when administrative costs are large relative to a product’s market. In a June 2026 progress report, Council delegations stressed the need to keep a balance between pre-market requirements and later safety monitoring. Administrative simplification does not, by itself, mean dropping the assessment of device safety.
The proposal also touches institutional cooperation, including support from the European Medicines Agency (EMA) for the expert panels on medical devices. It does not move the whole certification system to the EMA. The text under negotiation can still change substantially before any agreement between the Council and the European Parliament.
Health ministers took note of a progress report in June 2026; a Council general approach has not been adopted. The Irish Presidency of the Council has set itself the aim of a Council position by the end of 2026, but that remains a working timetable, not a decision. In the European Parliament, work on the file continues in the public-health committee. For the amendments to become law, the ordinary legislative procedure must be completed, involving both the Council and the Parliament.
Until any adoption, and until the dates of application set in a final act, the MDR and IVDR remain the applicable legal framework. Manufacturers and healthcare organisations should not treat the 11 September discussions as a change of the rules already decided. The negotiations still matter: how thousands of types of devices will be assessed, certified and monitored in the years ahead will affect industry, hospitals, laboratories, doctors and, ultimately, patients.
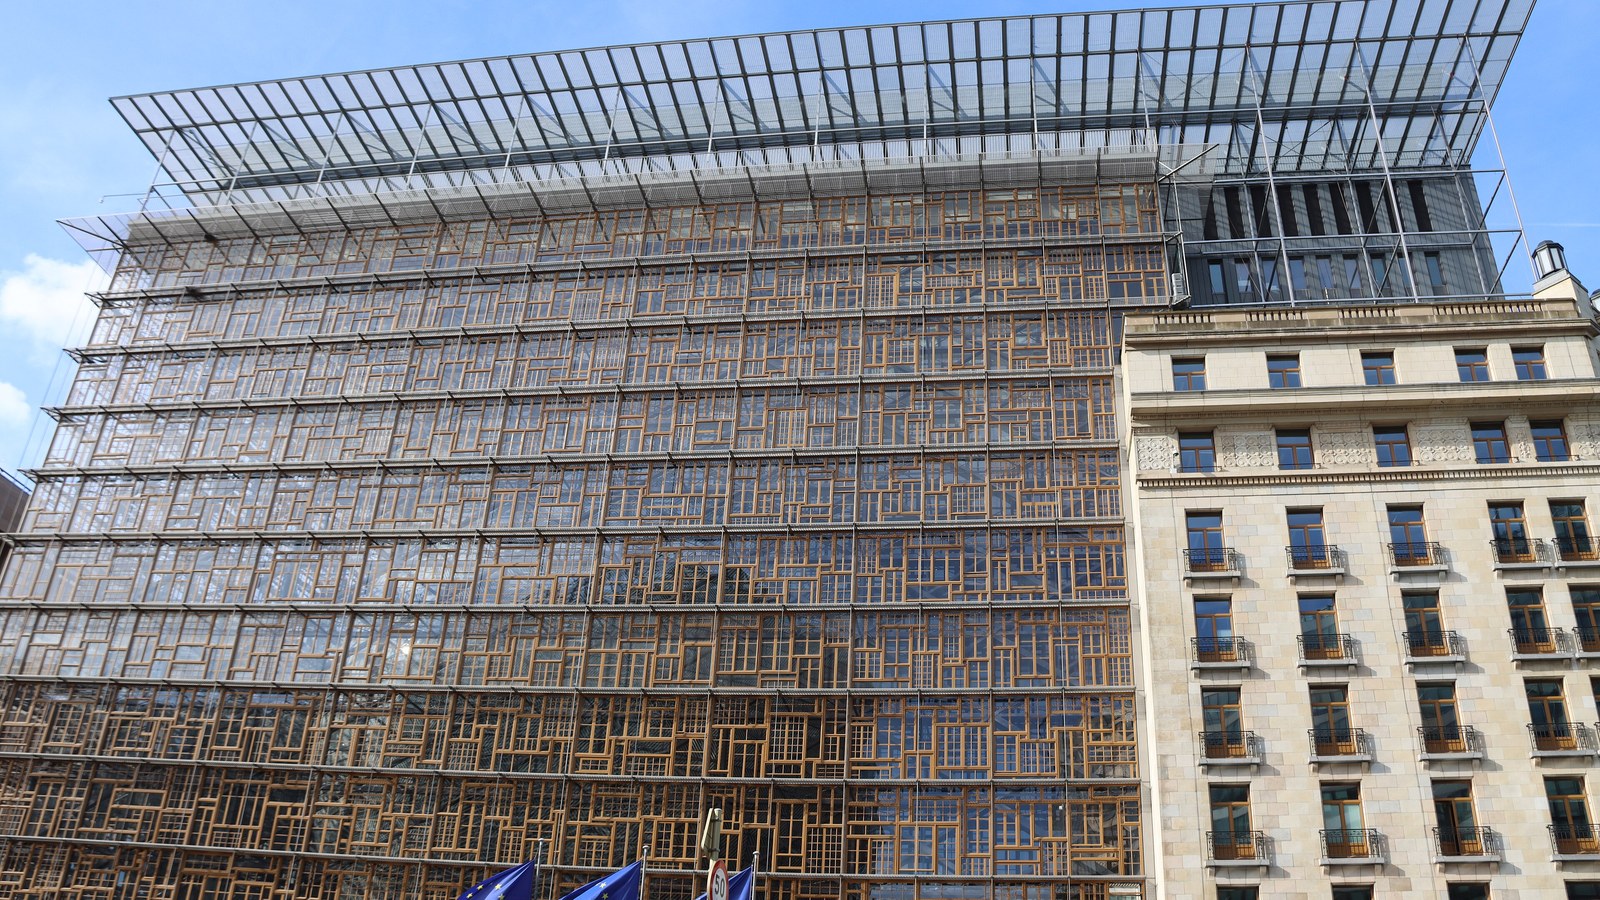
Photo: FrDr / Wikimedia Commons, CC BY-SA 4.0. The Europa building in Brussels, seat of the Council of the European Union, photographed on 1 July 2024. The image illustrates the institution, not the 11 September 2026 meeting. The photo was cropped to 16:9.
Sources consulted: COM(2025) 1023 — proposal for a regulation to simplify rules on medical devices and IVDs | EUR-Lex; Proposal for a regulation to simplify rules on medical and in vitro diagnostic devices | European Commission; Regulation to simplify rules on medical and in vitro diagnostic devices — Progress report | Council of the EU.
0 Comments
No comments on this article yet. Be the first!